Browse categories
Explore
Fiverr Pro
English
$
USD
Bioinformatics and Computational Chemistry, Vaccine Design, MD Simulation, DFT
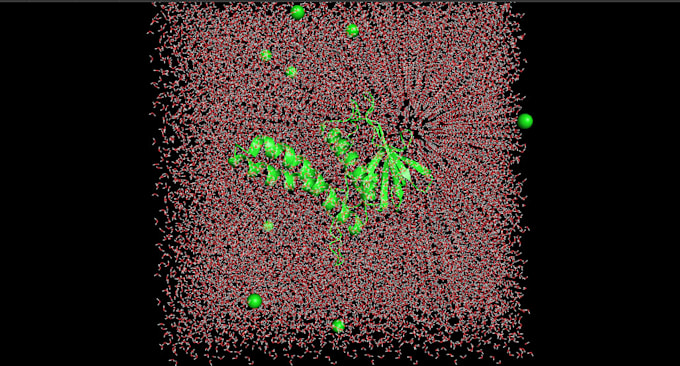
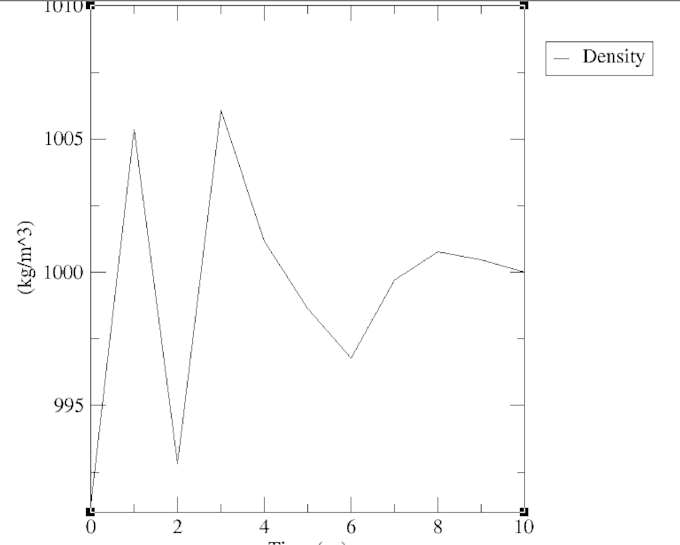
I run complete molecular dynamics simulations using GROMACS in a Linux environment energy minimization, NVT/NPT equilibration, production run, and post-simulation structural analysis with all results plotted and explained.
I can help you:
Ideal for students who need a full MD simulation with proper structural analysis for a thesis, assignment, or research project not just raw output, but a clear explanation of what it shows.


Technology:
Google Analytics
•
Google Data Studio
Analysis type:
Descriptive analysis
•
Predictive analysis
Expertise:
Prediction
•
Probability
•
Math
Programming language:
Python
What file formats do I need to provide?
A PDB file of your protein structure (or the raw sequence if you don't have one yet — I can help generate a structure first).
How long is the production simulation?
Standard runs are short simulations (suitable for initial stability checks). If you need longer production runs (multiple nanoseconds), message me before ordering so I can quote delivery time and pricing accurately.
What software do you use?
GROMACS on Linux, with analysis plots generated using Grace (xmgrace).
Will I understand the results if I'm not familiar with MD simulations?
Yes — every package includes a plain-language explanation of what the plots and analysis mean for your protein's stability, not just raw files.